The intricate balance of the human immune system, vital for survival, is designed to swiftly counteract threats such as infections or injuries before returning to a quiescent state. However, as individuals advance in age, this finely tuned response often goes awry, leading to a persistent, low-grade systemic inflammation known colloquially as "inflammaging." This chronic inflammatory state is increasingly recognized as a foundational contributor to a multitude of age-related pathologies, ranging from cardiovascular disease and neurodegeneration to metabolic disorders and certain cancers. Scientists have long sought to understand the molecular underpinnings of this phenomenon, particularly the role of cellular senescence – a state where cells cease dividing but remain metabolically active, often referred to as "zombie cells" due to their enduring presence and detrimental secretions.
In a significant stride toward deciphering this complex biological puzzle, a collaborative research endeavor involving institutions such as Sanford Burnham Prebys Medical Discovery Institute, the Mayo Clinic, and other key partners has unveiled a previously unrecognized, critical interplay between mitochondria, cellular metabolism, and the perpetuation of chronic inflammation. Their groundbreaking findings, meticulously detailed in the prestigious scientific journal Nature, delineate a dual mitochondrial signaling pathway that drives the inflammatory output of senescent cells. Furthermore, the study showcased a promising therapeutic intervention: targeted disruption of one component of this pathway successfully attenuated inflammation and supported improved healthspan in preclinical mouse models, illuminating a novel strategy for combating the deleterious effects of aging.
Cellular senescence, a state first described decades ago, typically arises when cells encounter various stressors, including irreparable DNA damage, telomere shortening, or oncogenic activation. Instead of undergoing programmed cell death (apoptosis) or continuing to divide with potentially harmful mutations, these cells enter a state of irreversible growth arrest. While this mechanism initially serves as a tumor-suppressive safeguard, preventing the proliferation of damaged cells, its accumulation with advancing age becomes problematic. Senescent cells are not inert; they actively reshape their local microenvironment through a complex array of secreted molecules known as the Senescence-Associated Secretory Phenotype (SASP). This SASP comprises pro-inflammatory cytokines, chemokines, growth factors, and proteases, collectively acting as potent drivers of inflammation, tissue dysfunction, and even the induction of senescence in neighboring healthy cells.
Dr. Peter Adams, a co-corresponding author on the study and the Jeanne and Gary Herberger Leadership Chair in Cancer Research at Sanford Burnham Prebys, underscored the dynamic nature of these cellular stalwarts. "Senescent cells are far from inactive," Adams elaborated. "They maintain high metabolic activity and possess an inherent inflammatory program that compels them to release a cascade of inflammatory mediators." It is this persistent secretion of SASP components that has been strongly implicated in the widespread chronic inflammation observed in older individuals, contributing significantly to the onset and progression of numerous age-related maladies.
The research team, spearheaded by the laboratory of Dr. João Passos, a senior and co-corresponding author and professor of Physiology at Mayo Clinic, embarked on a mission to dissect the precise molecular mechanisms that govern the activation and maintenance of the SASP. Their overarching objective was to identify potential vulnerabilities within these processes that could be therapeutically modulated or suppressed. Their investigations revealed a remarkable convergence of at least two distinct biological pathways, both intimately linked to the function and signaling capabilities of mitochondria – the ubiquitous organelles often dubbed the "powerhouses of the cell."
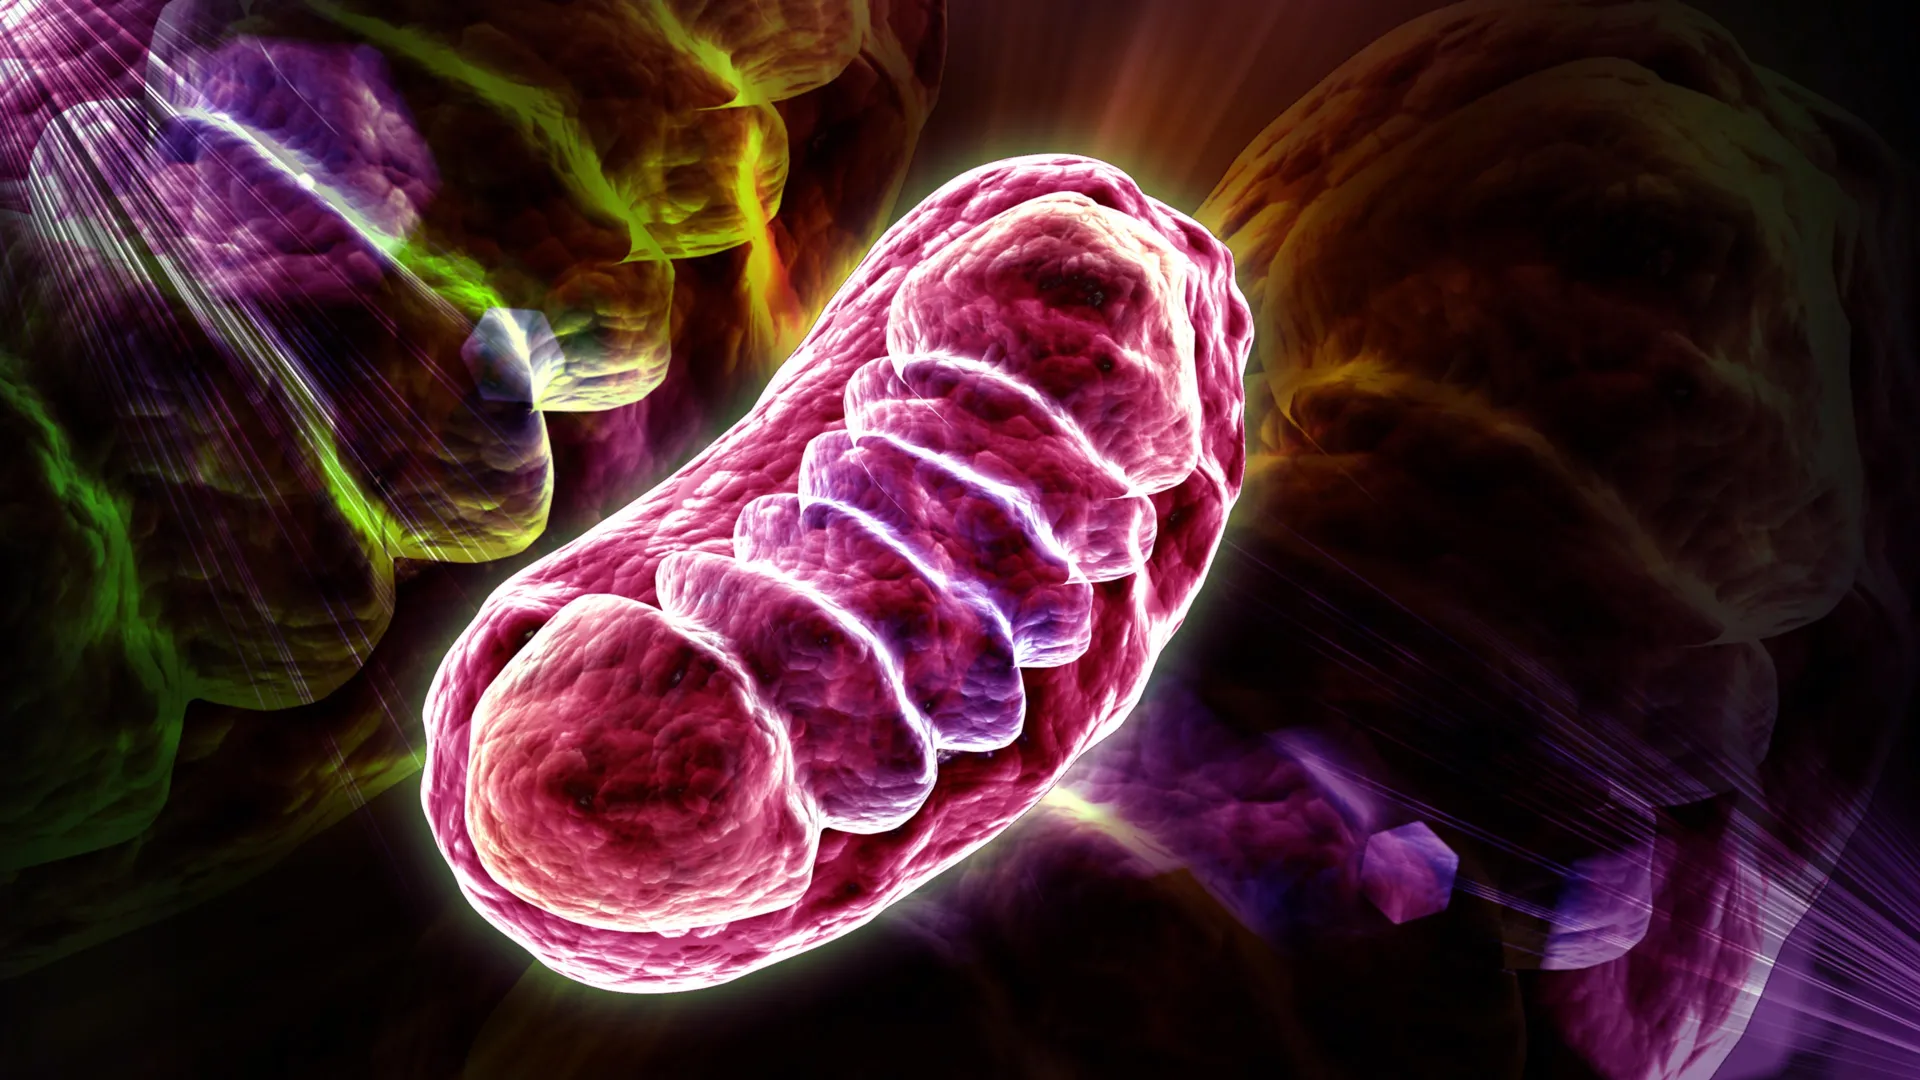
Mitochondria, beyond their primary role in generating adenosine triphosphate (ATP) through oxidative phosphorylation, are increasingly recognized as central hubs for cellular signaling, metabolic regulation, and immune responses. In senescent cells, the researchers observed a distinctive alteration in mitochondrial behavior compared to their counterparts in healthy, dividing cells. Specifically, mitochondria within senescent cells exhibited an elevated production of acetyl-CoA, a pivotal molecule that functions at the nexus of metabolism and epigenetics.
Acetyl-CoA’s influence extends beyond energy production; it plays a critical role in modifying histone proteins. Histones are spool-like proteins around which DNA is tightly wound to form chromatin, thereby regulating gene expression. By transferring an acetyl group to specific histone residues, acetyl-CoA triggers a process called histone acetylation. This modification effectively "loosens" the compact chromatin structure, decondensing the DNA and making certain genes more physically accessible to the cellular machinery responsible for transcription. Crucially, the researchers discovered that among these newly accessible genes were those encoding key components of the inflammatory SASP response. This metabolic reprogramming, therefore, acts as an epigenetic primer, setting the stage for subsequent gene activation.
However, the team’s meticulous analysis indicated that this metabolic shift and subsequent chromatin decondensation alone were insufficient to fully account for the continuous release of inflammatory molecules characteristic of senescent cells. A second, synergistic mitochondrial process was found to be indispensable.
Under conditions of cellular stress or damage, mitochondria themselves can become dysfunctional, leading to the leakage of their internal genetic material – mitochondrial DNA (mtDNA) and mitochondrial RNA (mtRNA) – into the cytoplasm, a compartment where these nucleic acids are not typically found. The cell’s innate immune system is exquisitely tuned to detect such misplaced genetic material, interpreting it as a "danger-associated molecular pattern" (DAMP). This detection triggers a robust inflammatory cascade, involving the activation of specific transcription factors. These transcription factors, once activated by the immune signals originating from leaky mitochondria, then target the very SASP genes that have already been rendered epigenetically accessible by the elevated acetyl-CoA levels.
In essence, the research unveiled a sophisticated two-pronged mechanism: one mitochondrial pathway, driven by altered metabolism and increased acetyl-CoA, prepares the inflammatory SASP genes by making them accessible through histone modifications. The second pathway, initiated by the release of mitochondrial genetic material, provides the critical immune signaling trigger necessary to actively switch on these primed SASP genes. "Witnessing how these two seemingly independent pathways intersect to promote SASP prompted us to investigate whether disrupting one could prevent their detrimental partnership," Dr. Adams remarked.
To test this compelling hypothesis, the researchers employed a pharmacological agent known as CTPI-2. This compound functions by inhibiting a specific transport protein crucial for supplying a precursor molecule required for acetyl-CoA synthesis. By blocking this transport, CTPI-2 effectively reduces the cellular production of acetyl-CoA, thereby attenuating the metabolic signal.
When CTPI-2 was administered to mouse models, the results were striking. The treatment led to a significant reduction in inflammation across various tissues throughout the animals’ bodies. Beyond merely dampening inflammatory markers, the intervention also yielded tangible physiological benefits, improving overall tissue function and extending the "healthspan" of the aging mice – a measure of the duration of healthy, functional life.
Importantly, the immune signals stemming from the leakage of mitochondrial DNA and RNA persisted even in the presence of CTPI-2. This indicated that the drug did not prevent mitochondrial damage or the subsequent immune alarm. However, by weakening the metabolic signal involving acetyl-CoA, the researchers effectively made the SASP genes less accessible for activation, thereby mitigating their inflammatory consequences. This observation highlights a crucial insight: while both pathways are necessary for full SASP expression, targeting the metabolic priming step alone is sufficient to disrupt the inflammatory cascade.
"Even with the continued immune signaling from compromised mitochondria, interfering with the metabolic signal rendered SASP genes less accessible, resulting in tangible functional improvements," stated Dr. Adams. "The strategic use of selective inhibitors, such as CTPI-2, to reduce acetyl-CoA and, consequently, inflammation, represents a novel and highly promising therapeutic approach that warrants extensive further investigation." He further emphasized the broader implications, suggesting that "this research fundamentally demonstrates that targeting metabolic signals that govern DNA accessibility could inaugurate a new paradigm for attenuating age-associated inflammation and the functional decline that accompanies it."
These findings represent a significant leap forward in understanding the fundamental mechanisms driving age-related chronic inflammation. By pinpointing the dual mitochondrial pathways that converge to amplify the SASP, the study not only provides profound insights into the biology of aging but also identifies new, actionable therapeutic targets. The potential to develop innovative pharmacological interventions, often termed "senomorphics," that modulate the behavior of senescent cells rather than simply eliminating them, holds immense promise for improving human healthspan and addressing the burgeoning public health challenge posed by age-related diseases. Future research will undoubtedly focus on translating these preclinical successes into safe and effective treatments for humans, potentially paving the way for healthier aging for generations to come.
The initial authorship of this pivotal publication is attributed to Hélène Martini, PharmD, PhD, a postdoctoral researcher within the Passos lab at Mayo Clinic. The comprehensive study received generous financial backing from multiple prestigious organizations, including the National Institutes of Health, National Institute on Aging, National Cancer Institute, National Institute of Diabetes and Digestive and Kidney Diseases, the Department of Defense Ovarian Cancer Research Program, the Hevolution Foundation, The Glenn Foundation for Medical Research, Cancer Research UK, and the Robert and Arlene Kogod Center on Aging.
About the Author